Anticorps Recombinant de lapin anti-SMN
SMN Recombinant Antibody for WB, IF/ICC, IP, ELISA
Hôte / Isotype
Lapin / IgG
Réactivité testée
Humain, rat, souris
Applications
WB, IF/ICC, IP, ELISA
Conjugaison
Non conjugué
CloneNo.
251620C4
N° de cat : 86668-2-RR
Synonymes
Galerie de données de validation


Applications testées
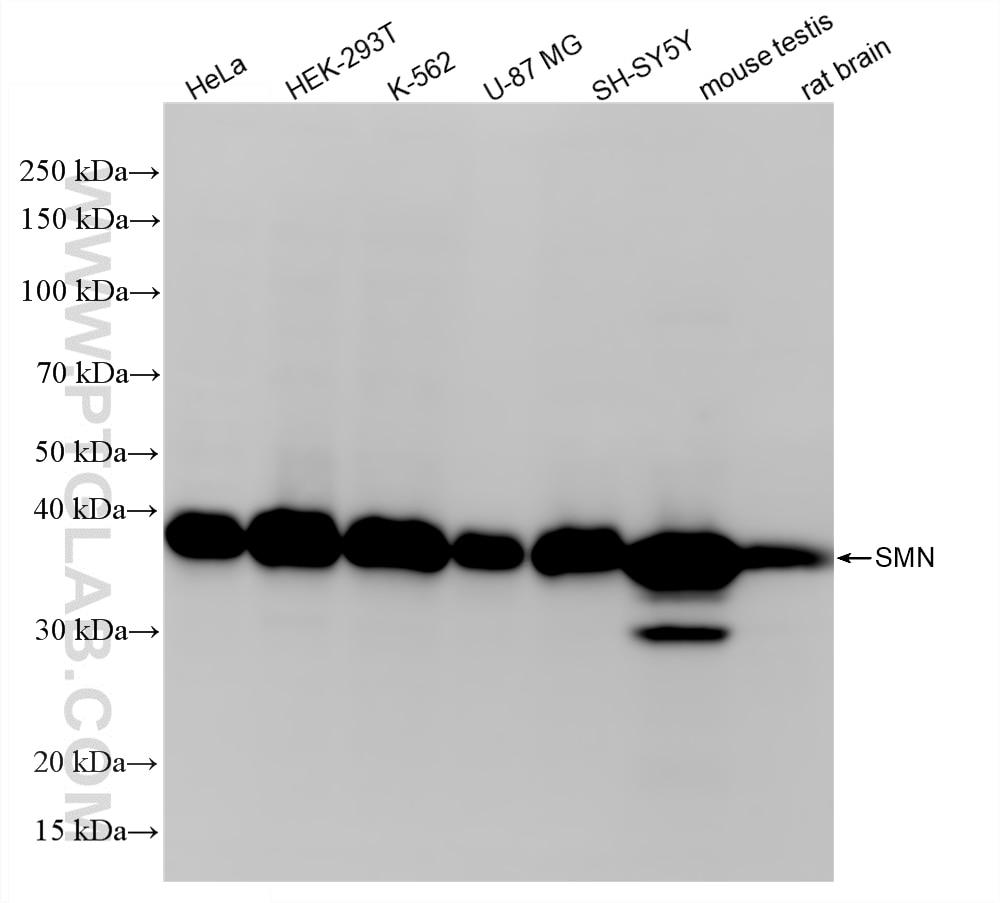
| Résultats positifs en WB | cellules HeLa, cellules HEK-293T, cellules K-562, cellules MG U-87, cellules SH-SY5Y, tissu cérébral de rat, tissu testiculaire de souris |
| Résultats positifs en IP | cellules HEK-293, |
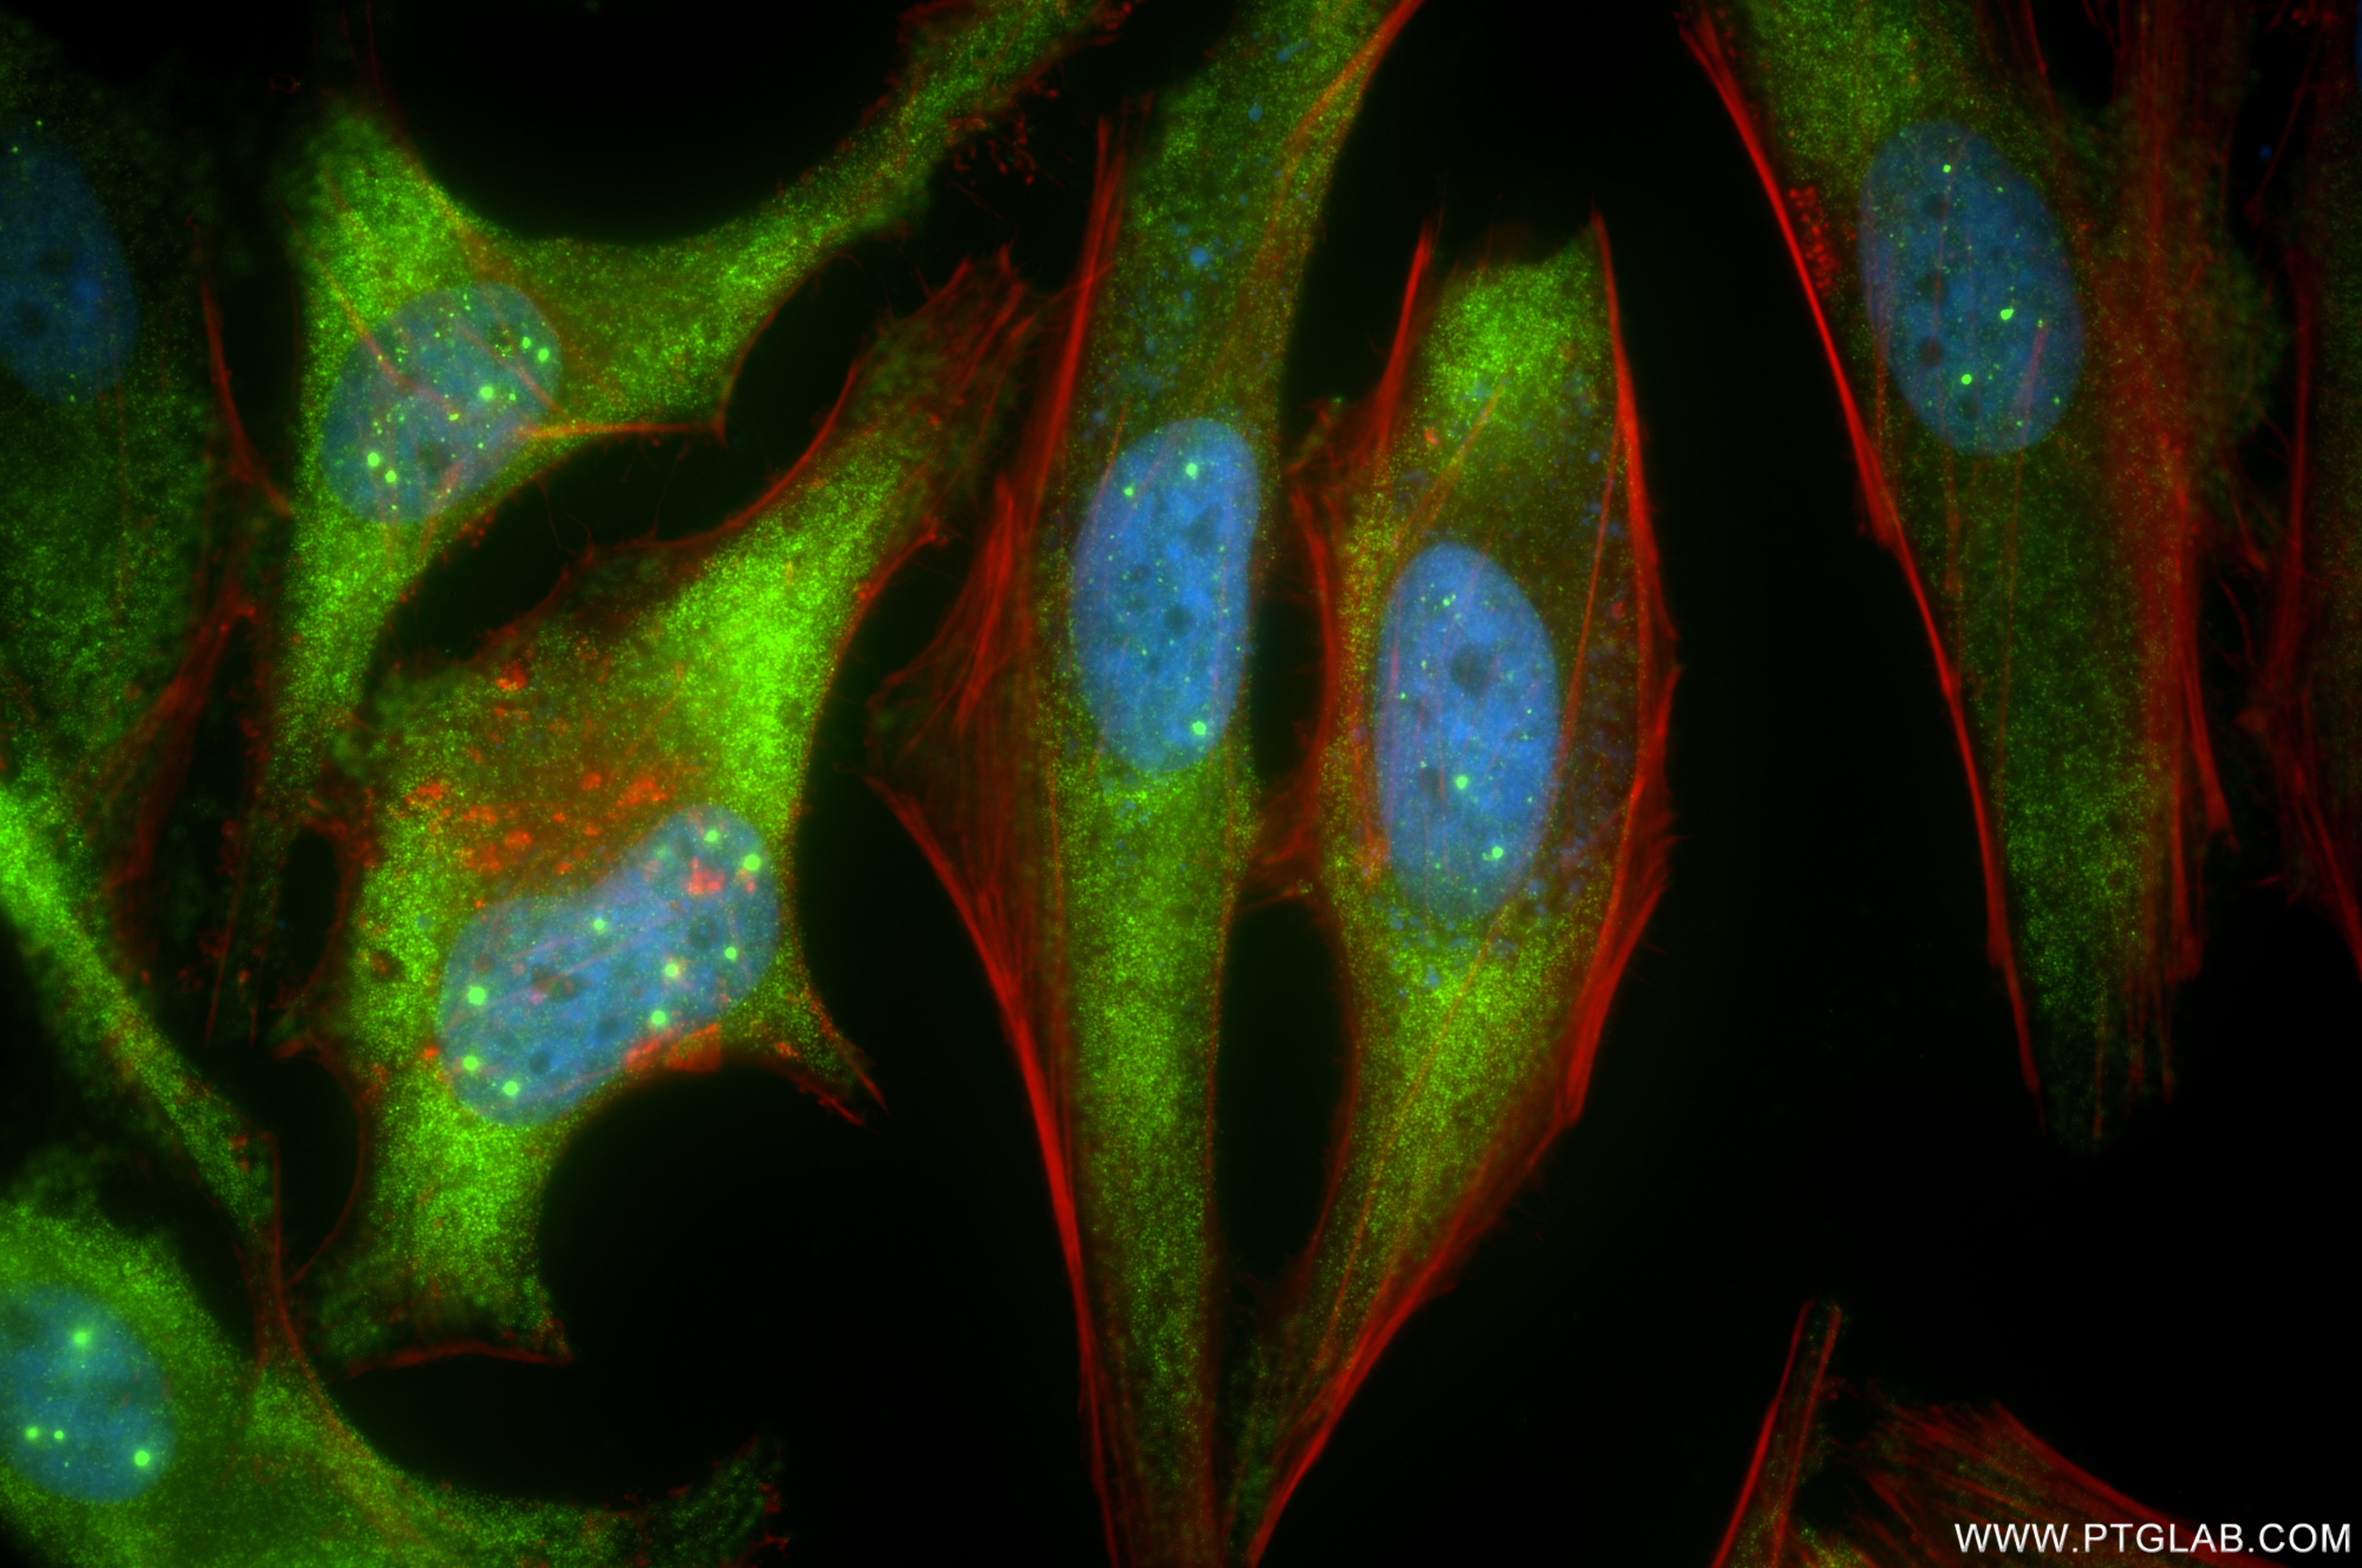
| Résultats positifs en IF/ICC | cellules HepG2, cellules HeLa |
Dilution recommandée
| Application | Dilution |
|---|---|
| Western Blot (WB) | WB : 1:1000-1:4000 |
| Immunoprécipitation (IP) | IP : 0.5-4.0 ug for 1.0-3.0 mg of total protein lysate |
| Immunofluorescence (IF)/ICC | IF/ICC : 1:250-1:1000 |
| It is recommended that this reagent should be titrated in each testing system to obtain optimal results. | |
| Sample-dependent, check data in validation data gallery | |
Informations sur le produit
86668-2-RR cible SMN dans les applications de WB, IF/ICC, IP, ELISA et montre une réactivité avec des échantillons Humain, rat, souris
| Réactivité | Humain, rat, souris |
| Hôte / Isotype | Lapin / IgG |
| Clonalité | Recombinant |
| Type | Anticorps |
| Immunogène | SMN Protéine recombinante Eg4812 |
| Nom complet | survival of motor neuron 2, centromeric |
| Masse moléculaire calculée | 30 kDa |
| Poids moléculaire observé | 38 kDa |
| Numéro d’acquisition GenBank | BC015308 |
| Symbole du gène | SMN |
| Identification du gène (NCBI) | 6607 |
| Conjugaison | Non conjugué |
| Forme | Liquide |
| Méthode de purification | Purification par protéine A |
| Tampon de stockage | PBS with 0.02% sodium azide and 50% glycerol |
| Conditions de stockage | Stocker à -20°C. Stable pendant un an après l'expédition. L'aliquotage n'est pas nécessaire pour le stockage à -20oC Les 20ul contiennent 0,1% de BSA. |
Informations générales
Spinal muscular atrophy (SMA) is an autosomal recessive neurodegenerative disease characterized by loss of anterior horn cells in the spinal cord and concomitant symmetrical muscle weakness and atrophy (PMID: 16364894 ). SMA is caused by deletion or mutations of the survival motor neuron (SMN1) gene. SMA patients lack a functional SMN1 gene, but they possess an intact SMN2 gene, which though nearly identical to SMN1, is only partially functional (PMID: 17355180). A large majority of SMN2 transcripts lack exon 7, resulting in production of a truncated, less stable SMN protein (PMID: 10369862). The level of SMN protein correlates with phenotypic severity of SMA.
Protocole
| Product Specific Protocols | |
|---|---|
| WB protocol for SMN antibody 86668-2-RR | Download protocol |
| IF protocol for SMN antibody 86668-2-RR | Download protocol |
| IP protocol for SMN antibody 86668-2-RR | Download protocol |
| Standard Protocols | |
|---|---|
| Click here to view our Standard Protocols |

